
ICH adopted M11 in November 2025 to fix the protocol format: what it covers, what it does not require, and where it changes the study build.
How an EDC system collects, checks and locks clinical trial data, what regulators expect from it, and the criteria that matter when you choose one.
What AI assistants and MCP are, how they help from protocol to closeout, and five principles for using them in regulated studies.

Regulation 2026/1744 pushes the AI Act's high-risk rules back to 2027 and 2028: what already applies to the AI in your trials, and what to prepare.
Forecasts, site stock levels, blinded kits and amendments: how investigational product moves from depot to participant, and what you must record at each step.
How academic and hospital research teams choose an EDC: grant budgets, no dedicated data manager, exports for the statistician, and the sponsor's own validation duty.
The EHDS timeline, article by article: what applies in 2027 and 2029, and why clinical trial data only enter secondary use in 2031.
Electronic information notices, accepted channels by content, multi-factor authentication, deadlines: what the CNIL's May 2026 update requires of your eConsent.
Risks, committees, statistics and MDR harmonization: what the fourth edition of ISO 14155 changes in your clinical investigation plan.
An adaptive trial changes during the study. For each of four complex designs, what your EDC must handle before you sign the contract.
Site retraining, edit check revalidation, legacy queries, RTSM updates and re-consent: the five costs an amendment adds after the eCRF change.
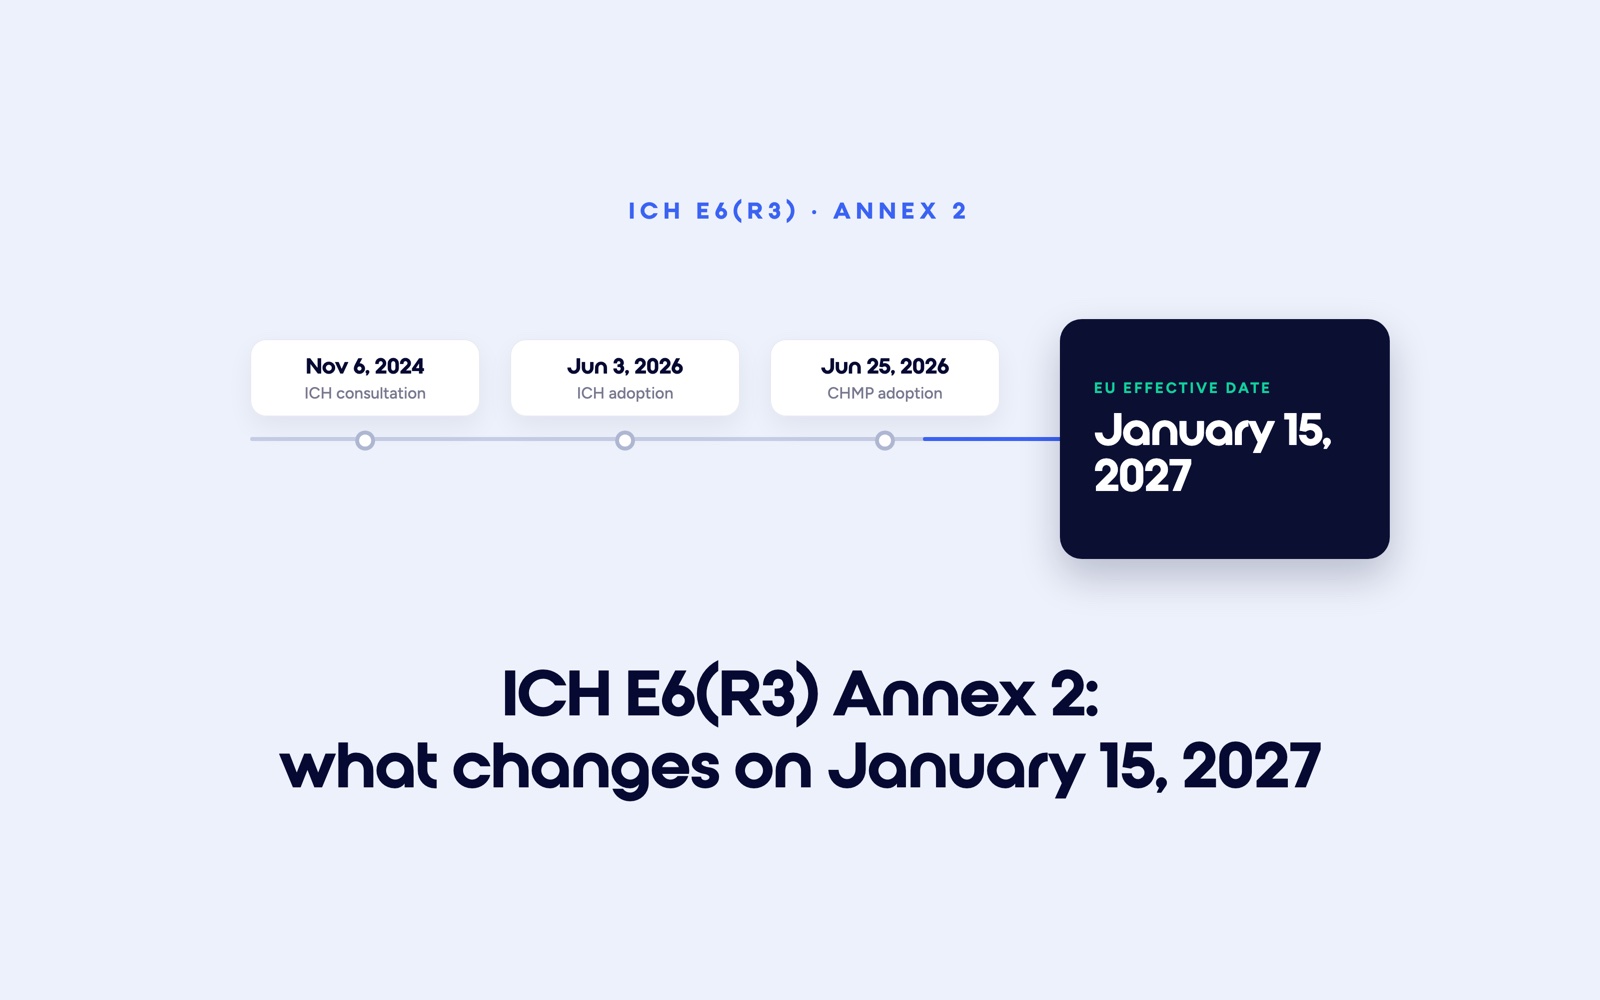
Protocol, remote consent and real-world data: what ICH E6(R3) Annex 2 asks of you before it takes effect in the EU on 15 January 2027.